
Highlights
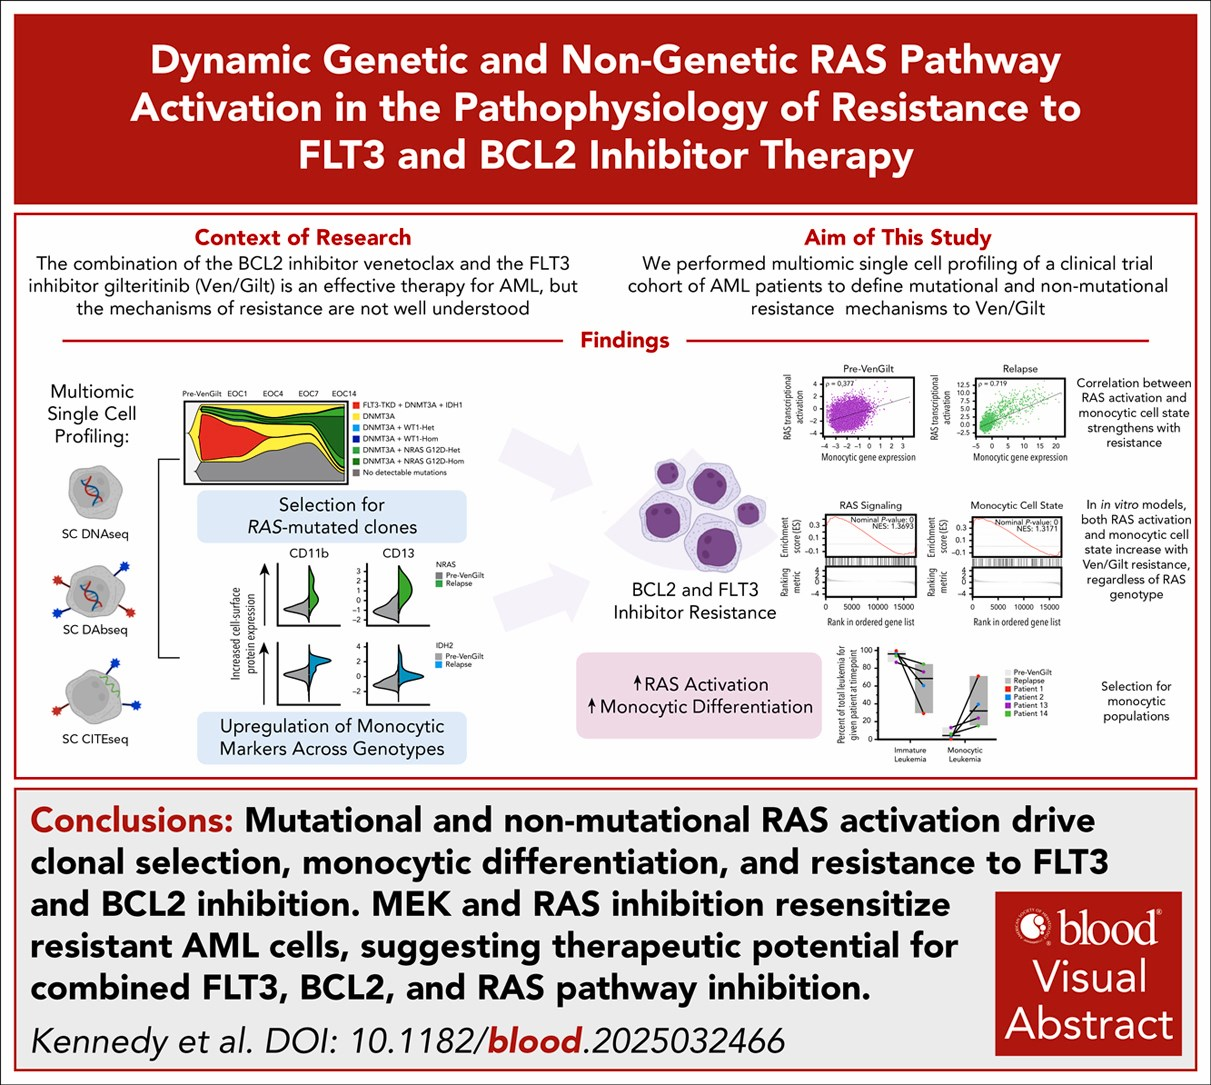
- Multiomic single-cell analyses reveal that resistance to combined FLT3 and BCL2 inhibitor therapy in AML is driven by diverse genetic (RAS mutations) and nongenetic (transcriptional activation) mechanisms activating the RAS pathway.
- Resistance involves a shift toward RAS-associated monocytic differentiation states, showing coupling between signaling pathways and cellular phenotype in AML relapse.
- FLT3-mutant clones are effectively eradicated by venetoclax and gilteritinib combination, but RAS-pathway activation emerges as a convergent resistance mechanism, underscoring the dynamic tumor evolution under treatment pressure.
- In vitro models demonstrate that RAS-pathway inhibition can resensitize resistant monocytic AML cells to Ven/Gilt therapy, suggesting potential therapeutic avenues to overcome resistance.
Background
Acute myeloid leukemia (AML) represents a heterogeneous hematologic malignancy characterized by clonal expansion of myeloid progenitors. Despite recent advances, relapse and resistance to targeted therapies remain formidable challenges. FLT3 mutations are prevalent in AML and confer poor prognosis. The BCL2 inhibitor venetoclax, often combined with hypomethylating agents, has improved outcomes but is hindered by resistance development. Similarly, FLT3 inhibitors like gilteritinib improve survival in FLT3-mutated AML but resistance pathways limit durable responses. Understanding mechanisms underlying resistance to dual FLT3 and BCL2 inhibition is critical to improving therapeutic strategies and clinical outcomes.
Key Content
Multimodal Single-Cell Profiling in Clinical Trial Patients
The paper by Kennedy et al. (Blood, 2026) used cutting-edge single-cell multiomic approaches combining DNA/protein and RNA/protein profiling on patient samples from a phase 1b clinical trial (NCT03625505) administering venetoclax plus gilteritinib (Ven/Gilt). This approach allowed high-resolution assessment of clonal architecture, transcriptional states, and surface immunophenotypes during therapy and at relapse, overcoming limitations of bulk sequencing.
Dynamic Clonal Evolution under FLT3 and BCL2 Inhibition
Ven/Gilt therapy robustly eliminated FLT3-mutant leukemic clones, confirming the efficacy of targeted FLT3 inhibition. However, resistance was marked by dynamic tumor evolution characterized by emergent clones harboring mutations in RAS-pathway genes, such as NRAS and KRAS, indicating genetic selection pressure. This supports prior evidence implicating RAS mutations as common resistance drivers in AML therapy (Papaemmanuil et al., NEJM 2016).
Nongenetic RAS Activation and Transcriptional Rewiring
Remarkably, the study uncovered nonmutational upregulation of RAS signaling transcriptional programs, independent of direct RAS mutations. This transcriptional activation was associated with AML cells adopting a monocytic differentiation phenotype, linked to RAS pathway activity. This aligns with previous observations that AML differentiation states are plastic and influence therapeutic resistance (van Galen et al., Cell 2019).
Monocytic Differentiation and Resistance Phenotype
AML cells transitioning towards monocytic lineage exhibit heightened RAS-pathway signaling and diminished sensitivity to apoptosis triggered by BCL2 inhibition. This phenotypic shift provides a cellular context in which resistance emerges despite effective FLT3 mutant clone eradication. Such differentiation-associated resistance adds complexity to target inhibition strategies.
Preclinical Validation of RAS-Pathway Inhibition
In vitro modeling of monocytic AML differentiation recapitulated the nonmutational activation of RAS pathways observed in patients. Pharmacologic inhibition of the RAS pathway re-sensitized these cells to venetoclax and gilteritinib, demonstrating translational potential for combinatorial targeting. This aligns with emerging preclinical data advocating for targeting downstream Ras/Raf/MEK/ERK signaling to overcome resistance (Zhang et al., Cancer Res 2021).
Broader Context and Evidence Synthesis
AML resistance mechanisms are multifactorial, involving clonal genetic evolution (e.g., FLT3, RAS mutations), epigenetic alterations, and transcriptional plasticity. The current study extends this paradigm by demonstrating that both genetic and epigenetic/nonmutational mechanisms can converge on RAS-pathway activation, a central hub for resistance. Notably, prior clinical studies have linked RAS mutations with resistance to FLT3 inhibitors (Smith et al., Blood 2020) and venetoclax (Pei et al., Leukemia 2020), but this work uniquely integrates multiomic single-cell data for higher resolution insights.
Expert Commentary
Kennedy et al.’s study is significant for several reasons. It employs state-of-the-art single-cell multiomics—a methodological advance providing unparalleled resolution of tumor heterogeneity and clonal dynamics. Their findings challenge the simplistic notion that resistance is solely mutation-driven, revealing that transcriptional and immunophenotypic shifts also drive therapeutic failure.
Clinically, these data argue for routine incorporation of approaches that capture not only mutational status but also cellular differentiation states and transcriptome remodeling to guide precision therapy. The demonstrated coupling of RAS activation with monocytic differentiation suggests that AML differentiation state is both a biomarker and therapeutic target.
The in vitro validation reinforces feasibility of RAS-pathway inhibitors in combination with FLT3 and BCL2 inhibitors, supporting development of clinical trials incorporating MEK or ERK inhibitors alongside Ven/Gilt. However, challenges remain, including identifying patients likely to benefit from triple therapy and managing added toxicity.
Importantly, the study’s insights may extend beyond AML to other cancers where RAS signaling and lineage plasticity contribute to targeted therapy resistance.
Limitations include the relatively small patient cohort size inherent to early-phase trials and the need for longer-term clinical outcome data. The complexity of clonal interplay and tumor microenvironment effects also warrant further investigation.
Conclusion
This comprehensive multiomic single-cell study reveals that resistance to combined FLT3 and BCL2 inhibition in AML involves dynamic genetic and nongenetic activation of the RAS pathway and is linked to monocytic differentiation shifts. These findings underscore the RAS pathway as a central node in therapeutic resistance and a compelling target for combination therapy.
This work highlights the necessity of integrated molecular and phenotypic profiling to understand and overcome resistance mechanisms, paving the way toward personalized combination therapies. Future clinical trials should explore the safety and efficacy of adding RAS-pathway inhibitors to current treatment regimens, aiming to improve durable remissions for AML patients.
References
- Kennedy VE et al. Dynamic genetic and nongenetic RAS-pathway activation drives resistance to FLT3 and BCL2 inhibitor therapy. Blood. 2026 Sep 17;148(12):1588-1604. PMID:42224378
- Papaemmanuil E et al. Genomic classification and prognosis in acute myeloid leukemia. N Engl J Med. 2016;374(23):2209-21. PMID:27135947
- van Galen P et al. Single-cell RNA-Seq reveals AML hierarchies relevant to disease progression and immunity. Cell. 2019 Apr 18;176(6):1265-1281.e24. PMID:30982665
- Smith CC et al. Clonal evolution and clinical resistance in sequential FLT3 inhibitor therapy. Blood. 2020;135(10):795-807. PMID:31824593
- Pei S et al. Targeting RAS signaling pathway to overcome resistance to venetoclax in AML. Leukemia. 2020;34(3):973-78. PMID:32050632
- Zhang J et al. MEK inhibition overcomes resistance in AML with activated RAS signaling. Cancer Res. 2021;81(17):4490-4501. PMID:34086545