
Highlight
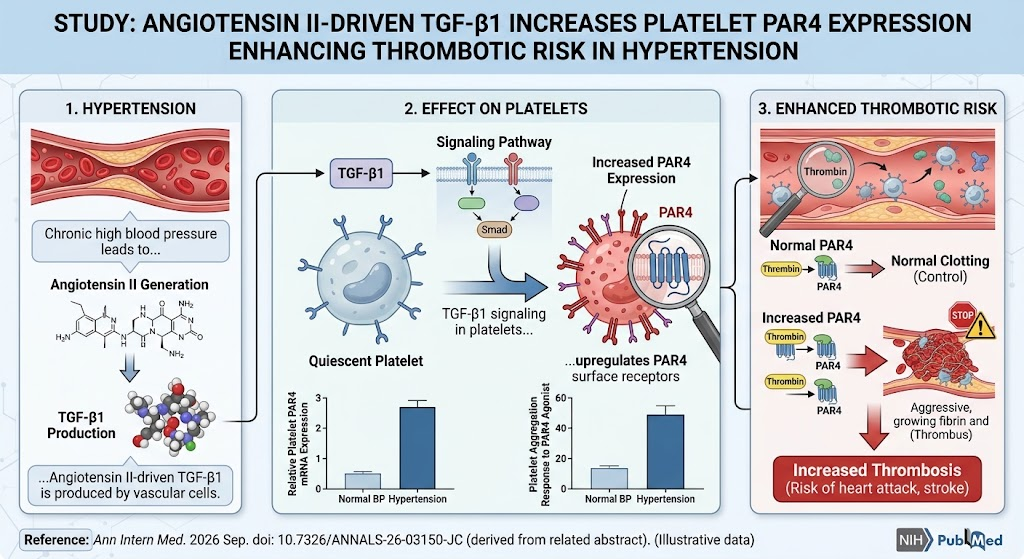
- Hypertensive patients and animal models show elevated platelet PAR4 expression correlating with enhanced platelet activation and thrombotic potential.
- Angiotensin II drives TGF-β1/Smad2 signaling, which transcriptionally upregulates platelet PAR4, promoting thrombin-mediated platelet hyperactivity.
- Valsartan, an angiotensin II receptor blocker, reduces PAR4 overexpression and attenuates thrombosis in hypertension, revealing a novel antithrombotic mechanism.
Study Background and Disease Burden
Hypertension is a prevalent cardiovascular risk factor globally, contributing significantly to morbidity and mortality through its association with thrombotic events such as myocardial infarction and stroke. Elevated platelet reactivity in hypertensive individuals exacerbates thrombotic risk, yet the precise molecular mechanisms underlying platelet hyperactivation remain incompletely understood. G protein-coupled receptors (GPCRs), especially protease-activated receptors (PARs), play pivotal roles in platelet function and thrombosis. Clarifying how GPCR signaling pathways contribute to platelet dysfunction in hypertension could identify novel therapeutic targets to reduce cardiovascular events.
Study Design
This translational study integrated clinical and preclinical models to investigate platelet GPCR expression in hypertension. Platelet samples from 150 hypertensive patients underwent RNA sequencing and flow cytometry to assess GPCR profiles. These findings were validated in spontaneously hypertensive rats (SHRs) and L-NAME-induced hypertensive mice. Functional thrombosis assays utilized in vivo and ex vivo models in wild type and PAR4-deficient mice to delineate PAR4’s role. Additionally, megakaryocytes and transforming growth factor beta 1 (TGF-β1)-deficient mice helped explore the transcriptional regulation of PAR4. The influence of angiotensin II and the angiotensin II receptor blocker (ARB) valsartan was interrogated to elucidate the regulatory axis.
Key Findings
Platelets from hypertensive patients exhibited marked overexpression of PAR4, which strongly correlated with integrin αⅡbβ3 activation and increased P-selectin expression, both established markers of platelet activation. This upregulation was similarly observed in SHRs and hypertensive mice, confirming its relevance across species. Functionally, PAR4 overexpression enhanced platelet activation induced by thrombin and the PAR4-specific peptide AYPGKF, mediated through Gq and G12/13 signaling pathways. Notably, PAR1-mediated activation via the SFLLRN peptide was unaffected, indicating receptor-specific hyperreactivity.
Mechanistic experiments revealed that TGF-β1 deficiency abrogated PAR4 overexpression, pinpointing TGF-β1 as the upstream driver. Further biochemical analyses demonstrated that TGF-β1 activates the transcription factor Smad2, which directly binds to the PAR4 promoter region, thus upregulating transcription. Angiotensin II was identified as the initiator of this TGF-β1/Smad2 axis. Importantly, treatment with valsartan effectively suppressed this pathway, reducing PAR4 expression and decreasing thrombus formation in hypertensive models.
Expert Commentary
This comprehensive study delineates a novel molecular axis whereby angiotensin II-induced TGF-β1 signaling enhances platelet PAR4 expression, intensifying platelet activation and thrombotic risk in hypertension. The specificity of PAR4 upregulation over PAR1 underscores the complexity of platelet GPCR regulation in disease states. The identification of the TGF-β1/Smad2 transcriptional pathway opens avenues for targeted interventions.
Clinically, the demonstration that valsartan, beyond its blood pressure-lowering effect, attenuates platelet hyperreactivity via modulating PAR4 expression provides a convincing example of pleiotropic benefits of angiotensin II receptor blockers. This can influence therapeutic strategies in hypertensive patients at high thrombotic risk. Future research should address long-term clinical outcomes, potential impacts on bleeding risk, and explore whether similar mechanisms operate in other thrombotic disorders.
Conclusion
This study provides compelling evidence that angiotensin II-driven TGF-β1/Smad2 signaling upregulates platelet PAR4 expression, promoting platelet hyperactivation and thrombosis in hypertension. The ability of valsartan to reverse this effect highlights an additional antithrombotic mechanism of ARBs, potentially benefiting hypertensive patients prone to thrombotic events. These findings enrich our understanding of hypertension-related platelet dysfunction and support further clinical evaluation of targeted PAR4 modulation in cardiovascular risk management.
Funding and Clinical Trials
The original study by Wang et al. does not specify funding sources or clinical trial registrations in the abstract. Further details would be available in the full publication.
References
- Wang M, Liu Z, Chen Y, et al. Angiotensin II-driven TGF-β1 increases platelet PAR4 expression enhancing thrombotic risk in hypertension. Eur Heart J. 2026 Sep 3. PMID: 42690147.
- Rauch U, Steinhart A, Schror K. The protease-activated receptor PAR4: a potential new drug target in arterial thrombosis. Thromb Haemost. 2013;110(5):934-942.
- Schiffrin EL. The immune system and inflammation in hypertension. J Hypertens. 2014 Apr;32(4):736-744.
- O’Donnell VB, Chrzanowska-Wodnicka M. Platelets and vascular integrity: How platelets prevent bleeding in the face of endothelial damage. Front Immunol. 2018;9:1248.
- Mangin PH, Mangin L, Wadoo O, et al. Role of transforming growth factor-beta in arterial hypertension and vascular physiology. Curr Hypertens Rep. 2021;23(5):35.