
Highlight
- Deletion of chromosome 17p [del(17p)] defines a high-risk subset in multiple myeloma (MM) with poor clinical outcomes.
- PKMYT1, a kinase involved in cell cycle regulation, is identified as a selective vulnerability in del(17p) or TP53-deficient MM cells.
- Pharmacological inhibition of PKMYT1 with RP-6306 induces DNA damage, mitotic catastrophe, and selective apoptosis in del(17p) MM cells.
- RP-6306 treatment reduces tumor burden and improves survival in preclinical del(17p)/TP53-deficient MM mouse models, supporting its potential as a biomarker-driven therapeutic option.
Background
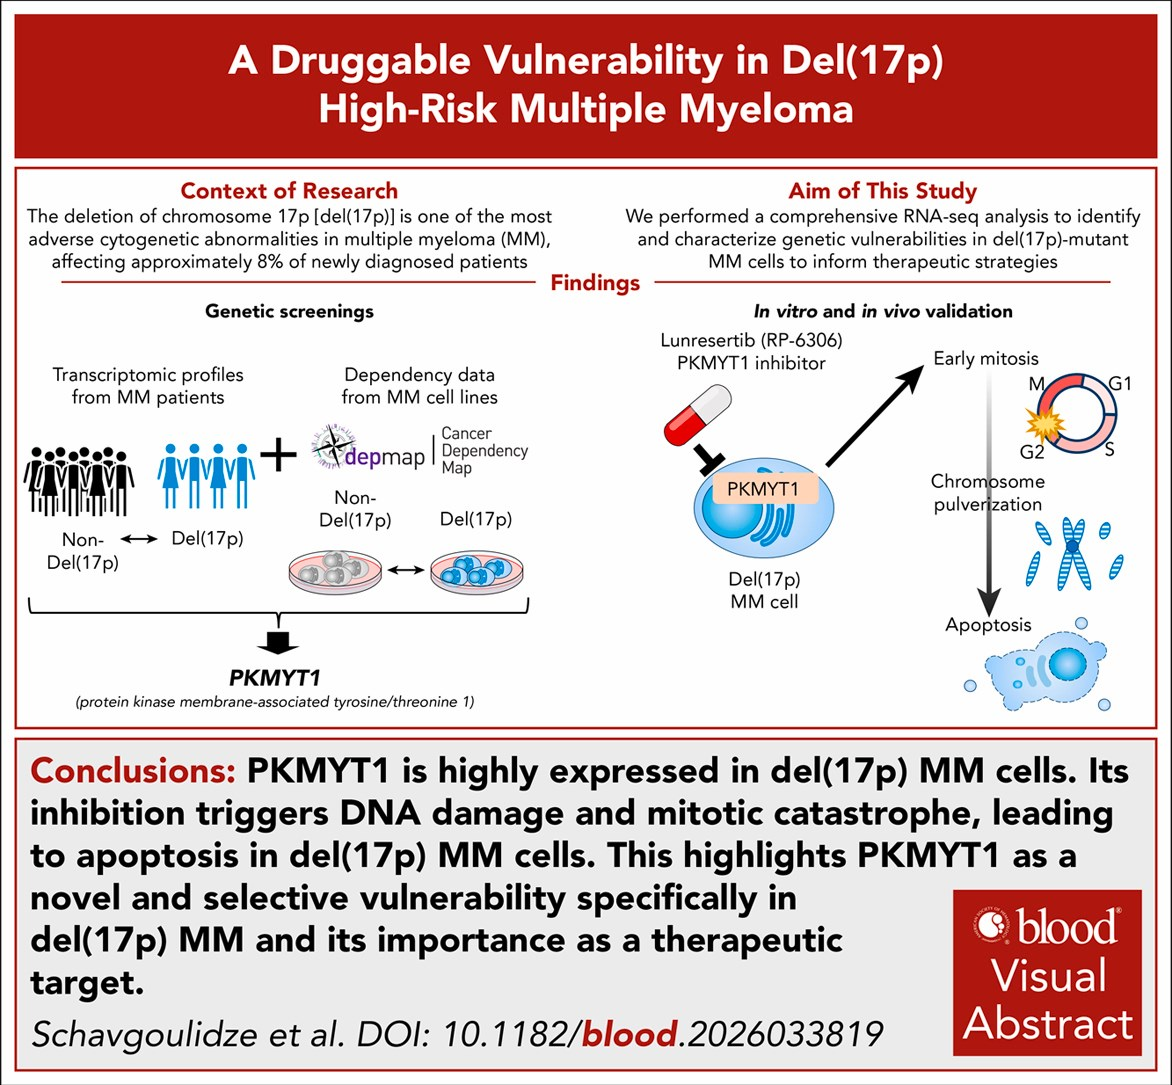
Multiple myeloma (MM) is a hematologic malignancy characterized by clonal proliferation of plasma cells within the bone marrow. Despite advances in treatment, MM remains incurable with heterogeneous outcomes largely influenced by cytogenetic abnormalities. Among these, deletion of the short arm of chromosome 17 [del(17p)], which encompasses the TP53 tumor suppressor gene, is strongly associated with high-risk disease, treatment resistance, and poor overall survival. Patients harboring del(17p) MM demonstrate an unmet medical need for novel, targeted therapies that can overcome intrinsic genomic instability and confer durable disease control.
Study Design
This translational research integrated RNA sequencing data from primary MM patient cells with genetic dependency screens of MM cell lines to identify novel therapeutic targets selectively essential in del(17p) MM. Focus was placed on kinases amenable to pharmacologic inhibition. PKMYT1, a WEE family kinase regulating cell cycle progression by inactivating CDK1, was nominated. In vitro genetic suppression via knockdown and pharmacological inhibition using RP-6306 were evaluated across MM cell lines with and without del(17p)/TP53-deficiency. Functional assays assessed DNA damage, micronucleus formation, cell cycle effects, and cell viability. In vivo therapeutic efficacy of RP-6306 was tested in xenograft models and TP53-deficient syngeneic mouse models. Key endpoints included MM cell death specificity, tumor burden reduction, and survival extension.
Key Findings
1. Selective Vulnerability of del(17p) MM Cells to PKMYT1 Inhibition
RNA-seq and dependency mapping revealed that PKMYT1 expression and activity were critical for survival of MM cells harboring del(17p) or TP53 mutations. Genetic knockdown of PKMYT1 selectively reduced viability of del(17p) MM lines compared to non-del(17p) and healthy cells.
2. Pharmacologic Targeting with RP-6306 Triggers DNA Damage and Mitotic Catastrophe
Treatment with the selective PKMYT1 inhibitor RP-6306 caused accumulation of DNA double-strand breaks, evidenced by increased gamma-H2AX foci, and micronucleus formation indicative of genomic instability. This disruption led to mitotic catastrophe, aberrant chromosome segregation, and apoptosis predominantly in del(17p) MM cells.
3. Favorable Therapeutic Window
Del(17p)-negative MM cells and normal hematopoietic cells were largely spared from RP-6306-induced cytotoxicity, supporting a selective therapeutic index.
4. In Vivo Anti-Myeloma Activity
RP-6306 administration decreased tumor burden and significantly prolonged survival in murine models of del(17p)/TP53-deficient MM. Both human MM xenografts and genetically engineered syngeneic models responded, demonstrating translational relevance.
5. Potential Biomarker-Driven Therapy
The dependency on PKMYT1 was closely linked to TP53 status, positioning PKMYT1 inhibition as a biomarker-driven strategy for patient selection and personalized therapy.
Expert Commentary
This study provides compelling mechanistic and preclinical evidence positioning PKMYT1 inhibition as an actionable vulnerability in the subset of MM patients with del(17p) and TP53 deficiencies. By exploiting cell cycle checkpoint dysregulation, PKMYT1 inhibitors drive lethal genomic instability selectively in tumor cells deprived of functional p53-mediated DNA damage response. The work offers a promising therapeutic avenue addressing a dire unmet need for high-risk MM. Limitations include the need for clinical trials to confirm safety and efficacy and to explore potential resistance mechanisms. Furthermore, broad applicability beyond del(17p) or combination with standard MM agents remains to be elucidated.
Conclusion
The identification of PKMYT1 as a selective targetable vulnerability in del(17p) MM advances our understanding of the molecular dependencies underpinning high-risk disease. Pharmacological PKMYT1 inhibition with agents like RP-6306 offers a rational, biomarker-guided treatment strategy disrupting critical cell cycle regulation and overcoming TP53-deficiency-associated treatment resistance. Ongoing clinical development could provide a novel precision medicine option improving outcomes for patients with aggressive MM.
Funding and Clinical Trials
The original study did not specify funding sources in the abstract. Further information can be obtained from the primary publication and clinical trial registries regarding ongoing evaluation of PKMYT1 inhibitors in MM.
References
1. Schavgoulidze A, Cui J, Mayoral JE, et al. PKMYT1 is a targetable vulnerability in del(17p) high-risk multiple myeloma. Blood. 2026;148(12):1559-1571. doi:10.1182/blood.2023020305.
2. Kumar SK, Rajkumar V, Kyle RA. Multiple myeloma. Nat Rev Dis Primers. 2017;3:17046.
3. Munshi NC, Anderson KC. Multiple myeloma: many roads to targeting a complex tumor. Nat Med. 2017;23(12):1439-1441.
4. Lecona E, Fernandez-Capetillo O. Targeting ATR in cancer. Nat Rev Cancer. 2018;18(9):586-595.
5. Bouwman P, Jonkers J. Molecular pathways: how can ATM deficiency sensitize tumors to PARP inhibitors? Clin Cancer Res. 2012;18(9):2767-2773.