
Highlight
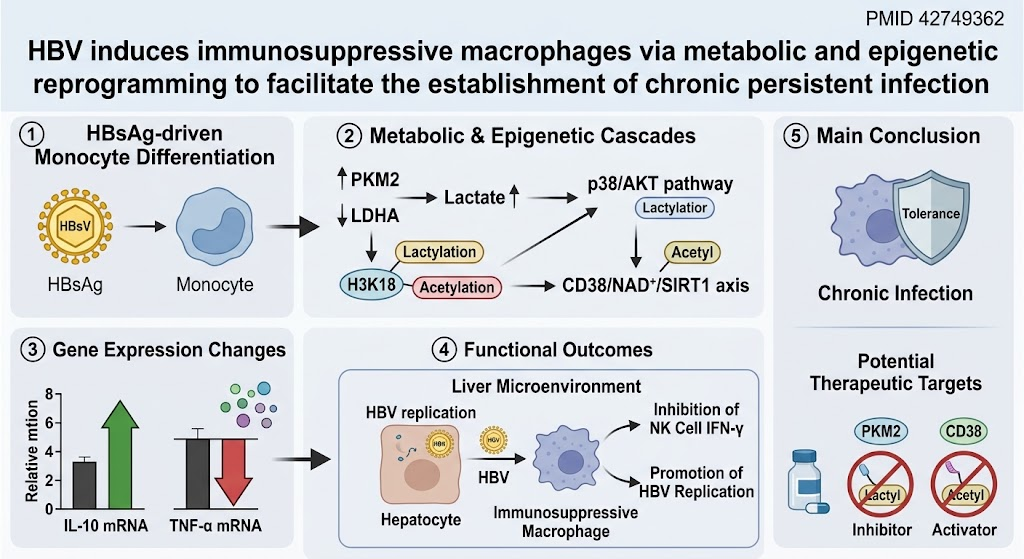
- HBV surface antigen (HBsAg) reprograms monocyte differentiation into immunosuppressive macrophages via metabolic and epigenetic mechanisms.
- HBsAg interacts with pyruvate kinase M2 (PKM2), promoting lactate production and H3K18 lactylation to enhance IL-10 expression, an anti-inflammatory cytokine.
- Altered STAT1 phosphorylation activates the CD38/NAD+/SIRT1 pathway reducing H3K18 acetylation at the TNF-α promoter, suppressing proinflammatory TNF-α production.
- HBV-educated macrophages inhibit NK cell IFN-γ secretion and facilitate HBV replication in hepatocytes, supporting viral persistence.
Study background
Chronic hepatitis B virus (HBV) infection remains a global health burden affecting over 250 million people worldwide and is a major cause of liver cirrhosis and hepatocellular carcinoma. A key factor in HBV persistence is the virus’s ability to evade host immune responses, particularly through induction of immunosuppressive environments within the liver. Macrophages play a central role in hepatic immunity and inflammatory regulation. Previous observations have noted the presence of immunosuppressive macrophages in chronic HBV infection, contributing to an environment that allows viral persistence. However, the detailed mechanisms by which HBV manipulates macrophage differentiation and function remain largely elusive.
Emerging research in infection immunology highlights that monocytes differentiate into macrophage subsets with distinct functional phenotypes influenced by metabolic and epigenetic reprogramming. Such processes have been documented in bacterial infections and after Bacillus Calmette-Guerin (BCG) vaccination, shaping either tolerogenic or trained macrophage responses. This study aimed to investigate whether HBV similarly exploits metabolic-epigenetic pathways during monocyte-to-macrophage differentiation to induce an immunosuppressive macrophage phenotype that facilitates chronic infection.
Study design
A translational experimental design was employed, analyzing monocytes from three sources: chronic HBV-infected patients, healthy adult controls, and human cord blood. Monocytes were differentiated in vitro into macrophages (monocyte-derived macrophages [MDMs] or cord blood monocyte-derived macrophages [CBMDMs]). The researchers characterized cytokine expression and secretion profiles, metabolic parameters, and epigenetic modifications using multiple methods including quantitative RT-PCR, ELISA, Western blotting, immunofluorescence, multiplex immunohistochemistry, immunoprecipitation, metabolite assays, ChIP-seq, ChIP-qPCR, plasmid transfection, and flow cytometry. Single-cell RNA sequencing data were reanalyzed for complementary insights. Functional assays examined the impact of HBV-educated macrophages on natural killer (NK) cell IFN-γ production and HBV replication in hepatocytes.
Key findings
Immunosuppressive macrophage phenotype in chronic HBV infection
Monocyte-derived macrophages from chronic HBV patients demonstrated an immunosuppressive phenotype characterized by elevated IL-10 production and reduced TNF-α secretion compared to macrophages from healthy donors. This cytokine balance favors immune tolerance over inflammation within the hepatic microenvironment.
HBsAg drives metabolic-epigenetic reprogramming in monocyte differentiation
Exposure to HBV surface antigen (HBsAg) notably influenced the differentiation of cord blood monocytes into macrophages exhibiting an immunosuppressive phenotype, whereas adult monocytes were less affected in vitro. Mechanistically, HBsAg binds and modulates pyruvate kinase M2 (PKM2), a key glycolytic enzyme, promoting the formation of PKM2 dimers which enhances PKM2/LDHA-mediated lactate production. Accumulating lactate mediates histone H3 lysine 18 lactylation (H3K18la) specifically at the IL-10 promoter region, epigenetically increasing IL-10 transcription and secretion.
Modulation of STAT1 phosphorylation and downstream signaling
HBsAg also altered STAT1 phosphorylation patterns by increasing serine 727 phosphorylation and decreasing tyrosine 701 phosphorylation via p38/AKT signaling. This shift activated the CD38/NAD+/SIRT1 axis, which decreased H3K18 acetylation at the TNF-α gene promoter, resulting in suppressed TNF-α synthesis. Thus, dual epigenetic modifications—lactylation promoting anti-inflammatory IL-10 and reduced acetylation limiting proinflammatory TNF-α—coordinate establishment of an immunosuppressive macrophage phenotype.
Functional consequences of HBV-educated macrophages
These macrophages inhibited natural killer (NK) cell production of IFN-γ, a critical antiviral cytokine, thereby impairing innate antiviral immunity. Moreover, co-culture experiments showed that HBV-infected hepatocytes had increased viral replication when exposed to HBV-educated macrophages. This indicates a feedback loop where HBV manipulates host innate cells to suppress antiviral responses and enhance viral persistence.
Expert commentary
This study provides compelling mechanistic insight into how HBV shapes the liver immune microenvironment to favor chronic infection. By linking viral antigen engagement with metabolic reprogramming and epigenetic histone modifications, it reveals a sophisticated evasion strategy targeting monocyte-to-macrophage differentiation. The identified pathways offer novel targets for therapeutic intervention aiming to restore effective immune responses against HBV. For example, inhibiting PKM2 dimerization or modulating histone lactylation/acetylation balance could potentially reverse macrophage immunosuppression.
Limitations include the in vitro nature of some experiments and the use of cord blood monocytes that may differ from adult cells in responsiveness. Further in vivo validation and clinical correlation are necessary. Nonetheless, this research aligns with growing recognition of metabolic-epigenetic axes as central coordinators of immune cell function in chronic infections.
Conclusion
HBV exploits metabolic and epigenetic reprogramming during monocyte differentiation through its surface antigen HBsAg to induce immunosuppressive macrophages. This mechanism suppresses proinflammatory responses and antiviral NK cell functions, creating a permissive environment for viral persistence and chronic infection. Targeting these pathways provides promising avenues for new therapeutics aiming to clear HBV and prevent liver disease progression.
Funding and clinicaltrials.gov
Detailed funding sources were not specified in the referenced publication. No registered clinical trials were indicated related to this mechanistic study.
References
1. Wang Z, Liu N, Li Y, et al. HBV induces immunosuppressive macrophages via metabolic and epigenetic reprogramming to facilitate the establishment of chronic persistent infection. Gut. 2026 Sep 16;PMID: 42749362.
2. O’Neill LA, Pearce EJ. Immunometabolism governs dendritic cell and macrophage function. J Exp Med. 2016;213(1):15-23.
3. Zhong Z, Sanchez-Lopez E, Karin M. Autophagy, Inflammation, and Immunity: A Troika Governing Cancer and Its Treatment. Cell. 2016;166(2):288-298.
4. Huang SC, Everts B, Ivanova Y, et al. Cell-intrinsic lysosomal lipolysis is essential for alternative activation of macrophages. Nat Immunol. 2014;15(9):846-855.